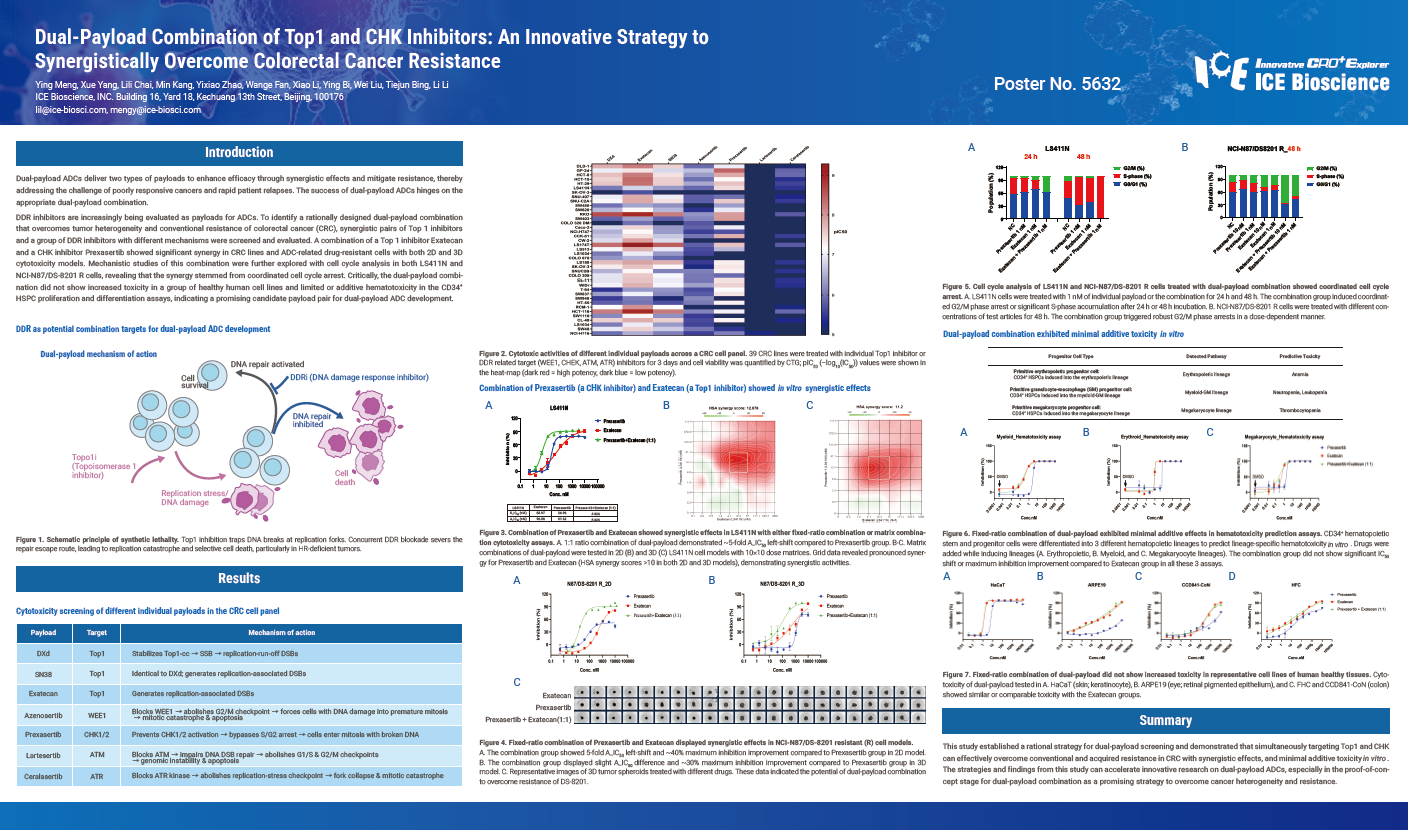
Dual-payload ADCs deliver two types of payloads to enhance efficacy through synergistic effects and mitigate resistance, thereby addressing the challenge of poorly responsive cancers and rapid patient relapses. The success of dual-payload ADCs hinges on the appropriate dual-payload combination.
DDR inhibitors are increasingly being evaluated as payloads for ADCs. To identify a rationally designed dual-payload combination that overcomes tumor heterogeneity and conventional resistance of colorectal cancer (CRC), synergistic pairs of Top 1 inhibitors and a group of DDR inhibitors with different mechanisms were screened and evaluated. A combination of a Top 1 inhibitor Exatecan and a CHK inhibitor Prexasertib showed significant synergy in CRC lines and ADC-related drug-resistant cells with both 2D and 3D cytotoxicity models. Mechanistic studies of this combination were further explored with cell cycle analysis in both LS411N and NCI-N87/DS-8201 R cells, revealing that the synergy stemmed from coordinated cell cycle arrest. Critically, the dual-payload combination did not show increased toxicity in a group of healthy human cell lines and limited or additive hematotoxicity in the CD34+ HSPC proliferation and differentiation assays, indicating a promising candidate payload pair for dual-payload ADC development.
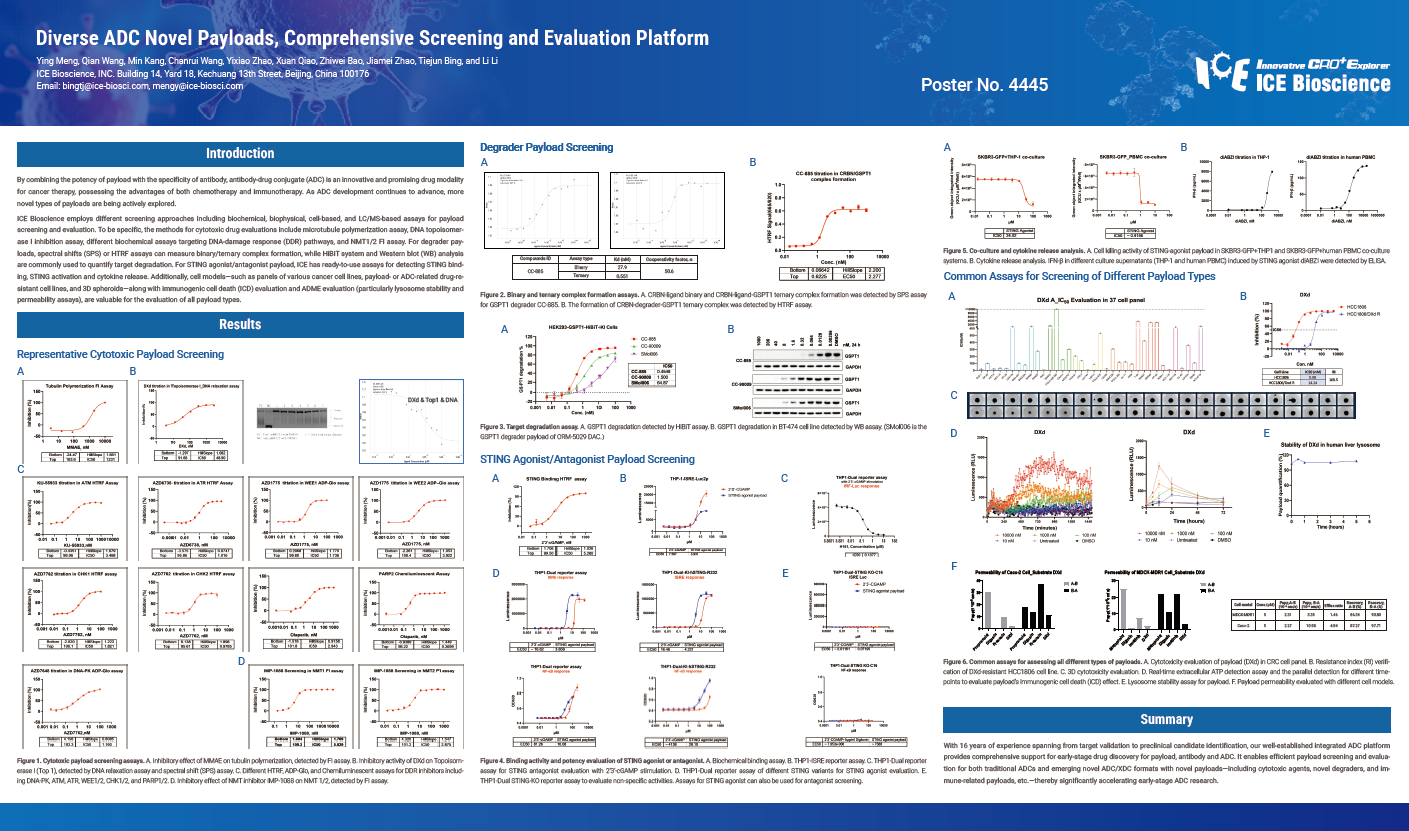
By combining the potency of payload with the specificity of antibody, antibody-drug conjugate (ADC) is an innovative and promising drug modality for cancer therapy, possessing the advantages of both chemotherapy and immunotherapy. As ADC development continues to advance, more novel types of payloads are being actively explored.
ICE Bioscience employs different screening approaches including biochemical, biophysical, cell-based, and LC/MS-based assays for payload screening and evaluation. To be specific, the methods for cytotoxic drug evaluations include microtubule polymerization assay, DNA topoisomerase I inhibition assay, different biochemical assays targeting DNA-damage response (DDR) pathways, and NMT1/2 FI assay. For degrader payloads, spectral shifts (SPS) or HTRF assays can measure binary/ternary complex formation, while HiBiT system and Western blot (WB) analysis are commonly used to quantify target degradation. For STING agonist/antagonist payload, ICE has ready-to-use assays for detecting STING binding, STING activation and cytokine release. Additionally, cell models—such as panels of various cancer cell lines, payload- or ADC-related drug-resistant cell lines, and 3D spheroids—along with immunogenic cell death (ICD) evaluation and ADME evaluation (particularly lysosome stability and permeability assays), are valuable for the evaluation of all payload types.

Breast cancer remains a leading cause of cancer-related mortality worldwide. While the simultaneous inhibition of PI3Kα and CDK4/6 has recently been clinically validated for advanced breast cancer, combination regimens of distinct targeted agents are often limited by overlapping toxicities and complex pharmacokinetics. To address these clinical hurdles and overcome therapeutic resistance, we developed a novel series of single-molecule dual PI3K/CDK4- 6 inhibitors by structure-based drug design strategy. Through extensive SAR studies, lead compounds 11 and 13 were identified, exhibiting nanomolar potency against PI3Kα and CDK4/6, high kinase selectivity, and potent antiproliferative activity in breast cancer cell lines. Compound 13 demonstrated robust in vivo efficacy in a T47D xenograft model, comparable to the Palbociclib–BKM120 combination with a good safety profile. This work highlights single- molecule dual PI3K/CDK4-6 inhibition as a highly promising therapeutic strategy for breast cancer.

We present an integrated hit discovery platform for B7‑H3 macrocyclic peptide binder screening that synergizes high‑throughput biophysical screening (SPR, SPS) with phage display technology. State-of-the-art protein structure prediction methods were employed to model the peptide–receptor complex, combined with binding free-energy calculations, thereby enabling the rapid identification and optimization of high-affinity cyclic peptide binders against oncology targets.
Our methodology employs parallel in vitro biophysical assays and phage display library screening. Protein structure prediction tools were also utilized to model the cyclic peptide–receptor complex structure, followed by binding free-energy calculations. The sequences obtained from phage display library screening were ranked and selected based on the Ipae metric and binding free-energy values. Target validation was achieved through engineered recombinant protein constructs (including B7-H3, FAP, EGFR, B7-H4 and GPC3) and isogenic cell lines with uniform target expression. Comprehensive evaluation was performed using modular functional assays measuring binding affinity and cellular internalization kinetics.
Address: Bldg 16, Yd 18, Kechuang 13th St, Etown, Tongzhou Dist, Beijing, 100176, China
Email: marketing@ice-biosci.com
Tel:+86-10-67809840